 K. Mark Wiencek
 02/22/2024

 Compounding Pharmacy,
USP <797>
Introduction
These standards include three key statements that we will discuss further in a two-part series:
-
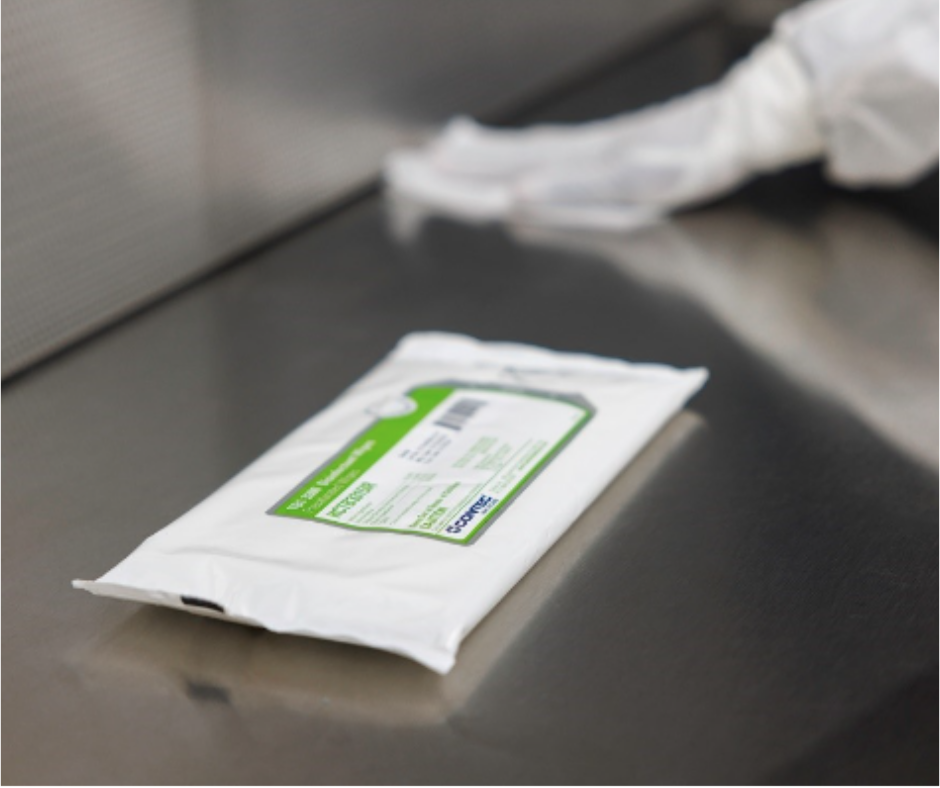
Cleaning and disinfecting agents used in the PEC (Primary Engineering Control) must be sterile.
-
Cleaning and disinfecting agents used in the SEC (Secondary Engineering Controls) or SCA should be sterile.
-
Sterile cleaning and disinfecting agents and supplies may be reused once opened, but only for a period specified by the manufacturer and/or described in the facility's written Standard Operating Procedures (SOPs).
Why is sterility important?
It is recognized there are many ways that microorganisms can enter spaces used for sterile compounding, including in the cleaning and disinfecting agents themselves. Certain forms of microbes, like bacterial spores, can survive in agents like 70% isopropanol (isopropyl alcohol, IPA), and could potentially survive inside any cleaning or disinfecting agent that is not sporicidal. While sporicidal agents pose a lower contamination risk, the packaging itself could harbor microbial bioburden and, therefore, should have sterile packaging. Finally, the requirement for sterile disinfectant agents aligns with the FDA’s Guidance for Industry titled “Insanitary Conditions at Compounding Facilities” (2020). If a cleaning agent were contaminated, the contamination would be spread across wide areas of the cleanroom and SCA during application. Even with the minimum surface sampling frequency increased to once per month, this widespread contamination could go undetected for several weeks.
Validated Sterile, Terminally Sterilized, SAL, Gamma Irradiated
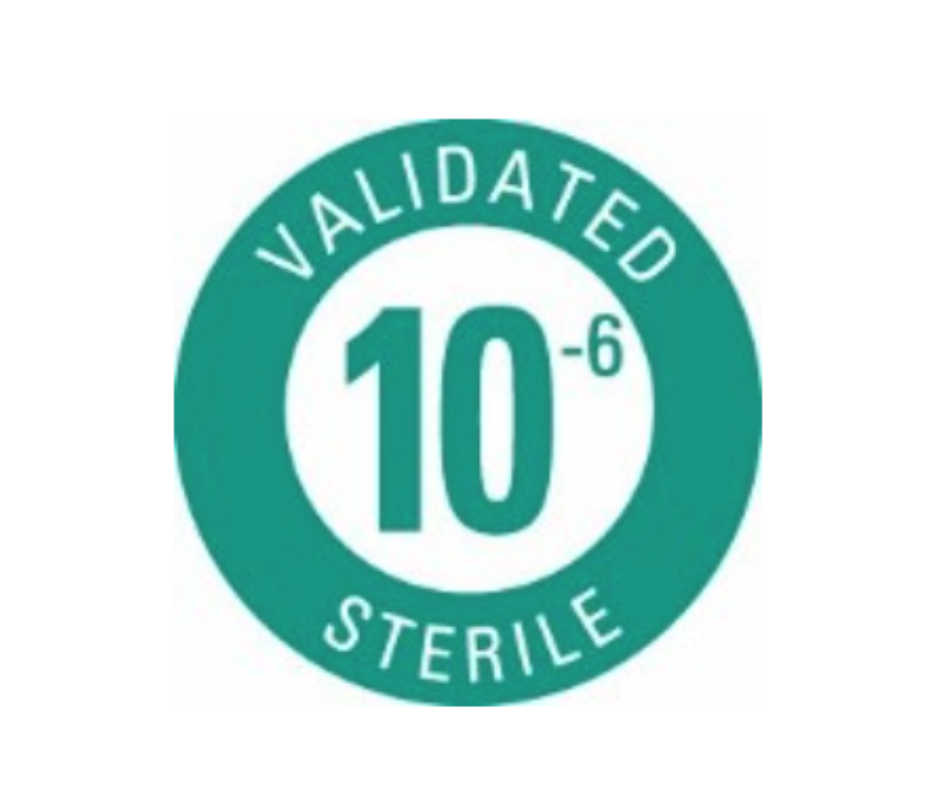
The Sterility Assurance Limit (SAL) is a term associated with the validation process, which refers to the probability that a product is not sterile. The most widely accepted SAL is 10-6, which means that no more than 1 out of 1 million (106) units may not be sterile. This validation process and SAL are necessary because it is impossible to test every unit for sterility.
Some consumable products, especially when first introduced to the market, may be irradiated to eliminate all microbes. Although this may reduce the risk of bioburden, simply irradiating the product alone does not make it terminally sterilized or “sterile.” Sterilization must be proven through validation efforts to applicable standards, and statistical analysis. If a product is only irradiated without this verification, it will not meet the criteria in the 2022 version of USP <797> or the FDA’s Guidance for Industry on Insanitary Conditions at Compounding Facilities
Additional Resources
 return to overview
 back to top